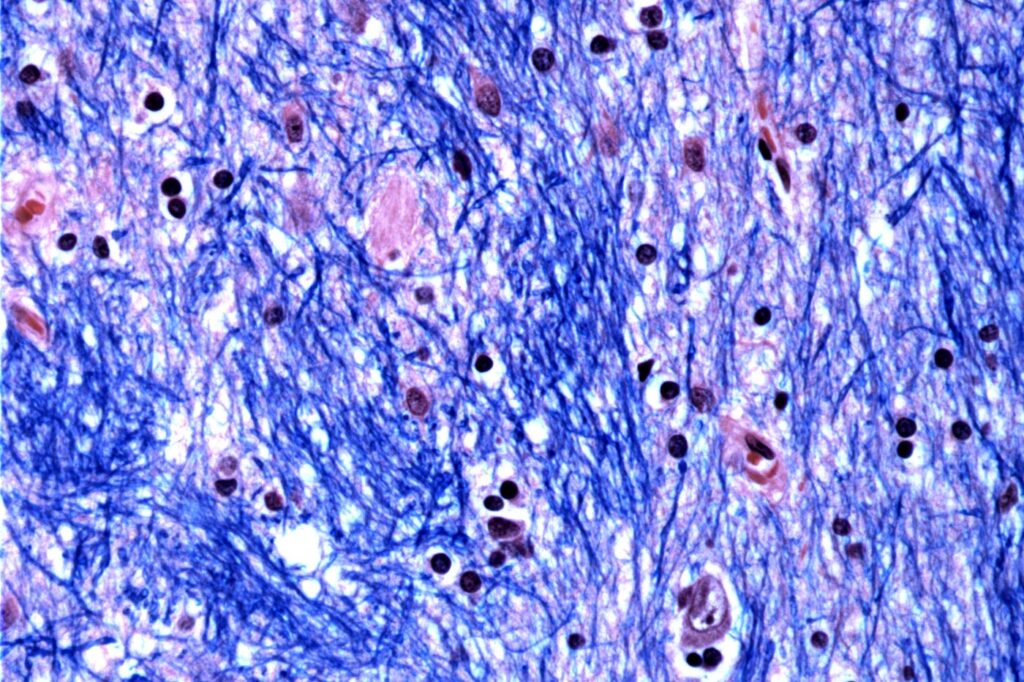
Huntington disease is a genetically inherited disorder that manifests with involuntary movements and psychiatric issues. While individuals with Huntington disease are born carrying the mutation, it remains unclear why some patients show symptoms much earlier than others. Researchers from the University of British Columbia's (UBC) Centre for Molecular Medicine and Therapeutics and the BC Children’s Hospital Research Institute aimed to unravel this mystery in their recent study.
Published in the scientific journal Neuron, the researchers discovered a crucial genetic variant that appears to accelerate the onset of Huntington disease by "driving runaway DNA changes" in the brain's most susceptible neurons. According to Dr. Michael Hayden, the senior author and a professor at UBC, individuals bearing this genetic variant experience a significantly hastened disease onset—by as much as 10 to 12 years—making the findings particularly impactful.
Dr. Hayden expressed relief in having identified the underlying process, stating, “This work answers that question and provides dramatic evidence that repeated expansion of the mutation is an important driver of Huntington disease and a potential treatment target.” The study delineates how the genetic variant contributes to a more rapid deterioration of neuronal health, which in turn precipitates earlier clinical symptoms of the disease.
The research team found that those with the genetic variant exhibited much larger expansions of the Huntington mutation in their neurons. This harmful expansion occurs approximately five times more frequently in patients with the variant compared to their counterparts without it. This heightened frequency of mutation expansion reinforces Huntington's classification as a primarily brain-centric disorder, even though the mutation affects every cell in the body.
These revelations culminate in a promising avenue for future therapeutic interventions. As stated by UBC researchers, the results may guide potential treatments targeting the mutation expansion process. The findings underscore the significance of repeat expansion of the DNA as a therapeutic target in Huntington disease. As Dr. Hayden pointed out, “If we can suppress that expansion, it may be possible to delay progression or delay the onset of disease.”
The implications of this study extend beyond mere academic interest; they offer hope for developing new strategies aimed at mitigating the effects of this devastating disease. By focusing on the molecular mechanisms that accelerate disease onset, the research paves the way for targeted therapies that could ultimately enhance the quality of life for those affected by Huntington disease.